Kuangbiao Liao, Solymar Negretti, Djamaladdin G. Musaev, John Bacsa & Huw M. L. Davies
Emory University, Atlanta, USA.
Nature 2016, 533, 230
http://www.nature.com/nature/journal/v533/n7602/pdf/nature17651.pdf
Abstruct
The laboratory synthesis of complex organic molecules relies heavily on the introduction and manipulation of functional groups, such as carbon–oxygen or carbon–halogen bonds; carbon–hydrogen bonds are far less reactive and harder to functionalize selectively. The idea of C–H functionalization, in which C–H bonds are modified at will instead of the functional groups, represents a paradigm shift in the standard logic of organic synthesis. For this approach to be generally useful, effective strategies for site-selective C–H functionalization need to be developed. The most practical solutions to the site-selectivity problem rely on either intramolecular reactions4 or the use of directing groups within the substrate. A challenging, but potentially more flexible approach, would be to use catalyst control to determine which site in a particular substrate would be functionalized. Here we describe the use of dirhodium catalysts to achieve highly site-selective, diastereoselective and enantioselective C–H functionalization of n-alkanes and terminally substituted n-alkyl compounds. The reactions proceed in high yield, and functional groups such as halides, silanes and esters are compatible with this chemistry. These studies demonstrate that high site selectivity is possible in C–H functionalization reactions without the need for a directing or anchoring group present in the molecule.

Strategy 2. Harnessing inherent character of substrate: White and co-workers Science 2010, 327, 566. (see also Science 2007, 318, 783.)


Davis group have been developed Rh2-dimer catalysts with kiral-carboxylate ligands.
See also "Hansen, J. & Davies, H. M. L. Highsymmetrydirhodium(II)paddlewheel complexes as chiral catalysts. Coord. Chem. Rev. 252, 545–555 (2008)."
Emory University, Atlanta, USA.
Nature 2016, 533, 230
http://www.nature.com/nature/journal/v533/n7602/pdf/nature17651.pdf
Abstruct
The laboratory synthesis of complex organic molecules relies heavily on the introduction and manipulation of functional groups, such as carbon–oxygen or carbon–halogen bonds; carbon–hydrogen bonds are far less reactive and harder to functionalize selectively. The idea of C–H functionalization, in which C–H bonds are modified at will instead of the functional groups, represents a paradigm shift in the standard logic of organic synthesis. For this approach to be generally useful, effective strategies for site-selective C–H functionalization need to be developed. The most practical solutions to the site-selectivity problem rely on either intramolecular reactions4 or the use of directing groups within the substrate. A challenging, but potentially more flexible approach, would be to use catalyst control to determine which site in a particular substrate would be functionalized. Here we describe the use of dirhodium catalysts to achieve highly site-selective, diastereoselective and enantioselective C–H functionalization of n-alkanes and terminally substituted n-alkyl compounds. The reactions proceed in high yield, and functional groups such as halides, silanes and esters are compatible with this chemistry. These studies demonstrate that high site selectivity is possible in C–H functionalization reactions without the need for a directing or anchoring group present in the molecule.
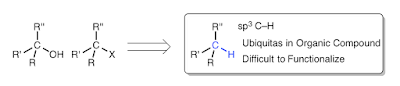
Impact of direct functionalization of alkane C–H bonds

How to achieve selective functionalization of C–H?
Strategy 1. Using direction group: Daugulis and co-workers J. Am. Chem. Soc. 2005, 127, 13154.
(Direction group: Murai*, Kakiuch, Chatani et al. Nature 1993, 366, 529)
Strategy 2. Harnessing inherent character of substrate: White and co-workers Science 2010, 327, 566. (see also Science 2007, 318, 783.)

Electronic and stereo electronic effects based on presence of heteroatom.
Strategy 3 (This work). Construction of finely designed pocket

(15 examples)


Reactive species (maybe Rh-carben species) is set on the bottom of the finely designed asymmetric cavity with enough depth, D2 symmetry, and rigidity.

1. Site Selective (0 : 25 : 1), 2. Diastereo Selective (20 : 1), 3. Enantio Selective (99%ee)
See also "Hansen, J. & Davies, H. M. L. Highsymmetrydirhodium(II)paddlewheel complexes as chiral catalysts. Coord. Chem. Rev. 252, 545–555 (2008)."
コメント